An inspector opens your CAPA log, picks a record that was closed six months ago, and asks you to walk through it from the first line to the last. That single question tells us more about a quality system than an hour of small talk. A closed CAPA nobody can explain end to end (what triggered it, why it happened, what changed, how you know it worked) is a finding waiting to be written.
We have sat through enough of these walkthroughs, and re-opened enough "closed" CAPAs, to know that the process written in the SOP and the process that actually survives scrutiny are frequently two different things. This is what the second one looks like, in a plant running under EDA or FDA-aligned GMP.
What Actually Triggers a CAPA
A CAPA does not start with a form. It starts with a signal, and the quality of your program depends on how many signal sources actually feed into it. The obvious ones are deviations and out-of-specification results. The ones plants routinely under-use are customer complaints, internal and external audit observations, trend data from routine testing, change control risk assessments, equipment maintenance logs, and near misses that never became a documented deviation because nothing was technically out of spec that day.
If your CAPA log is fed almost entirely by deviations, that is itself worth noticing. It usually means complaints are handled in a separate silo, trending is done in a spreadsheet nobody reviews quarterly, and audit findings get a one-line response instead of a proper investigation. An inspector who samples your complaint file and finds a pattern your CAPA system never caught will ask why the two systems do not talk to each other.
Writing a Problem Statement an Inspector Can Follow
Most weak CAPAs are weak from the first paragraph. "Tablet hardness was out of range" is an observation, not a problem statement. A problem statement needs to answer what happened, where, when, how it was detected, what the immediate impact was, and how it differs from the expected or specified condition. Vague scoping here is what later lets an investigation wander into an unfocused root cause, or worse, converge on the first plausible explanation instead of the real one.
We tell QA teams to write the problem statement as if handing it to someone who was not in the room: a new hire, or an auditor six months later. If that reader would still have questions about what actually happened, the statement is not finished. This single habit fixes more downstream CAPA quality problems than any tool or software change.
Root Cause: 5 Whys, Fishbone, and Knowing Which One to Reach For
Root cause is where most of the real work happens, and where "operator error" gets written far too often as a lazy stand-in for a systemic gap. We covered the mechanics of these tools in depth in our root cause analysis guide; here is how to pick between them for a CAPA specifically.
5 Whys for contained, single-cause events
The 5 Whys method works well when a deviation has one clear causal chain: a machine setting, a missed step, a specification gap. The failure mode most Egyptian plants fall into is stopping at two or three whys, landing on "the operator did not follow the SOP," and closing the investigation there. The real question is always why the SOP allowed that failure to happen in the first place — a missing in-process control, an ambiguous instruction, a training gap that was never closed out.
Fishbone for multi-factor or recurring events
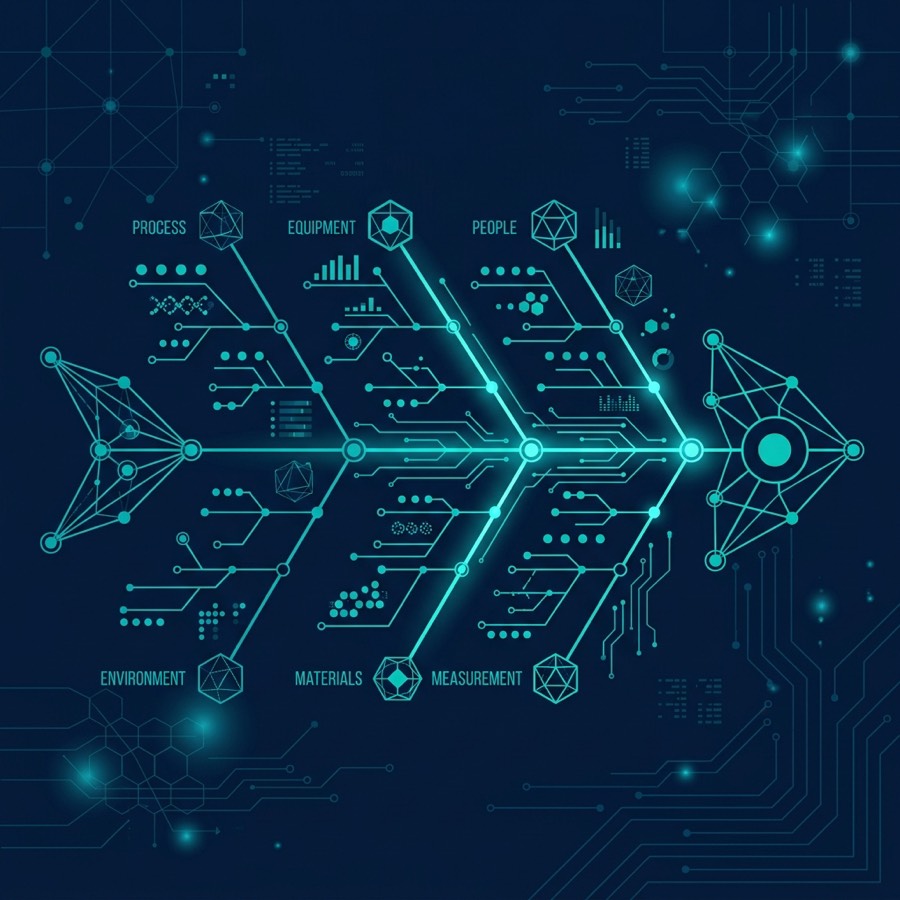
When a problem has touched more than one category of cause (man, machine, method, material, measurement, environment), a fishbone session run with the actual people involved (operators, not just supervisors) usually surfaces a cause nobody would have guessed from a desk. This is also the right tool when the same deviation type has recurred two or three times and each prior investigation reached a different, unrelated root cause. That pattern itself is a signal the earlier investigations were too shallow.
Corrective Actions vs Preventive Actions: Not the Same Line Item
A correction addresses what is in front of you right now: reject the batch, requarantine the material, recalibrate the instrument. A corrective action changes the process, procedure, equipment, or training so the identified root cause cannot reproduce the same failure. A preventive action addresses a weakness found before it caused a nonconformance — from a risk assessment, a trend, or a near miss on a different line that shares the same equipment or procedure.
We still see CAPA records in Egyptian plants that fold all three into one action line, usually something like "retrain operator and update SOP." That single line cannot be verified for effectiveness, because it is really three different claims bundled together. Separate them, assign an owner and a due date to each, and effectiveness checks stop being guesswork.
Effectiveness Checks: Where Most CAPAs Quietly Fail
This is the step regulators scrutinize hardest, because it is the step most quality teams skip or fake. Closing a CAPA because the retraining session happened and the SOP revision was approved is not effectiveness evidence — it is completion evidence. Effectiveness evidence has to look forward: has the specific failure mode recurred during a defined monitoring window? Do trend charts for the related parameter stay within control limits across the next several batches? Does a targeted audit of the changed step, three months later, still find it being followed correctly?
A useful discipline is to define the effectiveness check and its monitoring period at the same time you approve the corrective action, not after the action is already implemented. If nobody wrote down what "effective" would look like in advance, closure becomes a judgment call made under deadline pressure, and that is exactly the pattern an inspector is trained to notice.
Common CAPA Failures Inspectors Flag
Across regulated industries, the citation patterns are boringly repetitive, which is useful because it tells you exactly where to look before someone else does:
- Root cause recorded as "human error" with no supporting investigation of why the system allowed the error to occur.
- CAPAs closed on completion of the action, with no effectiveness data collected afterward.
- The same failure mode recurring across multiple CAPAs that were each treated as an isolated, unrelated event.
- Due dates extended repeatedly without documented justification or management visibility into the delay.
- A corrective action that does not logically address the stated root cause — the two paragraphs simply do not connect.
- No linkage between a CAPA and the change control, training record, or document revision it actually required.
None of these are missing features. They are gaps in discipline that a paper-based or loosely-enforced electronic system lets slide, because nothing forces the record to be complete before it can be closed.
How an eQMS Enforces the CAPA Loop
This is the part software actually earns its place. A properly configured CAPA software module does not just store the record, it enforces the sequence: a CAPA cannot move to "closed" until root cause is documented, until each corrective and preventive action has an owner and a due date, and until an effectiveness check with its own monitoring period has been recorded against the record. Extending a due date requires a reason and an approval, so the audit trail shows who decided to extend it and why, instead of a due date that silently slipped.
The other place an eQMS pays for itself is linkage. A CAPA rarely lives alone: it usually traces back to a record in deviation management, and it should trace forward into training records, document revisions, or change control. When those modules sit inside the same QMS platform instead of separate spreadsheets and shared drives, an auditor can follow one thread from the original deviation through root cause, through the corrective action, to the effectiveness data, without you assembling a binder overnight before the inspection. That traceability, more than any single feature, is what turns "we investigated it" into evidence an inspector can actually verify.
Frequently Asked Questions
What is the difference between a correction, a corrective action, and a preventive action?
A correction fixes the immediate problem in front of you, such as rejecting a bad batch or re-cleaning a line. A corrective action changes the process, procedure, training, or design so the same root cause cannot produce the same failure again. A preventive action addresses a weakness identified before it has caused a nonconformance, often from a trend, a risk assessment, or a near miss. CAPA records should capture all three separately rather than folding them into one vague action line.
How long should a CAPA investigation take?
There is no universal regulatory number, but most quality systems set internal targets by severity: a few days for a minor deviation-driven CAPA, and several weeks for a critical one involving product quality or patient safety. The timeline should be defined in your CAPA procedure and tracked, because an inspector will ask why a record sat open for months without documented justification or an approved extension.
What counts as CAPA effectiveness evidence?
Effectiveness evidence is data collected after the action was implemented that shows the specific failure mode has not recurred over a defined monitoring period: repeat deviation rates, trend charts, requalification results, or targeted audit findings. A signature confirming the action was completed is not effectiveness evidence. The check has to look at outcomes, not at whether a task was closed on time.
Can a CAPA stay open if the root cause is a business decision, like budget for new equipment?
Yes, but it should stay open honestly rather than being closed on paper. Interim controls should be documented and justified while the permanent action is pending, and the record should show management is aware of the risk being carried. An inspector is far less concerned by an honestly-open CAPA with interim controls than by one closed early to make a metric look better.
Do all deviations need a CAPA?
No. Many deviations are one-off events resolved with a correction and a documented root cause that does not point to a systemic gap. A CAPA is warranted when the root cause is process-level, when the same failure mode has recurred, when severity or risk to product quality is high, or when a trend across multiple minor events reveals a shared underlying cause. The decision criteria should be written into your deviation and CAPA procedures, not made case by case from memory.
Want CAPA records that hold up under inspection?
See how CORPEX-CAPA enforces root cause, effectiveness checks, and full traceability back to the originating deviation.
Explore CORPEX-CAPA